
A Deeper Dive into the MHRA Good Pharmacovigilance Practice (GPvP) Inspection Metrics and Findings
A Deeper Dive into the MHRA Good Pharmacovigilance Practice (GPvP) Inspection Metrics and Findings
Regulatory Affairs Vigilance Pharma & Biotech
Thursday, 28 March 2024

Anam Ahmad, Pharmacovigilance Consultant at QbD Group

 Share
 \
\
X (Twitter)
 \
\
LinkedIn
 \
\
WhatsApp
 \
\
Copy link
 \
\
Mail
Gain an insightful understanding into how the MHRA inspect the Pharmacovigilance services of an MAH for post-authorised products.
Play
A Deeper Dive into the MHRA Good Pharmacovigilance Practice (GPvP) Inspection Metrics and Findings | QbD Group
AI-generated audio
6:52
The UK Medicines and Healthcare Products Regulatory Agency (MHRA) has been conducting Good Pharmacovigilance Practice (GPvP) inspections since 2003. Any Marketing Authorisation Holder (MAH) or marketing authorization applicant is subject to a GPvP inspection to determine their compliance with the necessary pharmacovigilance duties required, in order to monitor the safety of medicines to the public in the United Kingdom (UK).
Inspections are typically scheduled by Pharmacovigilance System, product or post-authorisation safety study, rather than by MAH. To establish safety and compliance, there must be a Pharmacovigilance System in place for each MAH which describes in detail, the legal roles and responsibilities required to demonstrate the safety of the authorised medicinal product, in accordance with any changes to the risk-benefit ratio of the product.
The MHRA GPvP Symposium was held on 28th February 2024, where a range of topics were discussed to give an insight into how the MHRA achieved their goal of safeguarding the health of the public.
In this blog, the focus is more on the inspection metrics, to highlight the key areas where MAHs can continue to improve their PV system and prevent those critical, major or minor findings!
Types of Inspections
The objective of routine PV inspections is to assess whether the MAH can identify, characterise and report new or changed risks associated with their medicinal products.
Routine inspections can be either initial or re-inspections and are scheduled as part of the MHRA’s national inspection program according to a risk-based approach in line with the Good Pharmacovigilance (GVP) Module III. The MHRA will provide notice to the MAH before the inspection for preparedness.
Depending on the findings, the MAH may be subject to a re-inspection, otherwise, if there are no critical findings, there is no specific timeline by which an MAH must be inspected again, and this will be determined using a risk based approach.
Triggered inspections, also known as ‘ for cause inspections’ can be scheduled due to intelligence or a previous critical finding. This includes if the MHRA have been informed of specific risk information regarding the MAH and its authorised products. The following breaches of the GPvP also call for a triggered inspection:
-
a whistleblower
-
other MHRA departments
-
another regulatory authority
For these types of inspections, very little or no notice is given in advance.
MHRA GPvP Inspection Metrics for 2022/2023
There were 17 PV inspections conducted in 2022/2023 which included routine inspections and triggered inspections.
A significant number of inspections conducted during this period were ‘for cause – intelligence’ inspections. The graph below shows the breakdown of findings issued by the MHRA along with the inspection type.
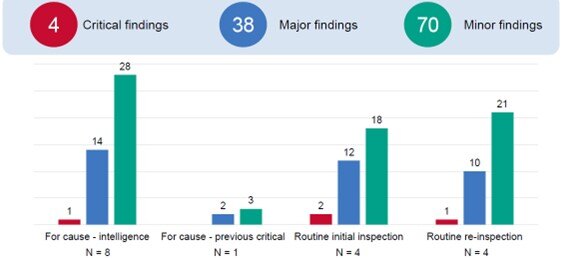 Figure 1: Findings based on Inspection Type
Source: Medicines and Healthcare products Regulatory Agency
The majority of critical findings were issued for routine initial inspections, followed by routine re-inspection and ‘for cause – intelligence’ inspections. The major and minor findings for ‘for cause – previous critical’ inspections are significantly lower than other inspection types, which is likely due to the limited scope for these types of inspections.
A robust Quality Management System (QMS) is required within a Pharmacovigilance system for the MAH. Though, the QMS may not be inspected directly by the MHRA, the majority of the major and minor findings were found in this area due to the QMS underpinning the surrounding PV topics. Two critical findings were issued for Risk Management and one for Ongoing Safety Evaluation which is illustrated below.
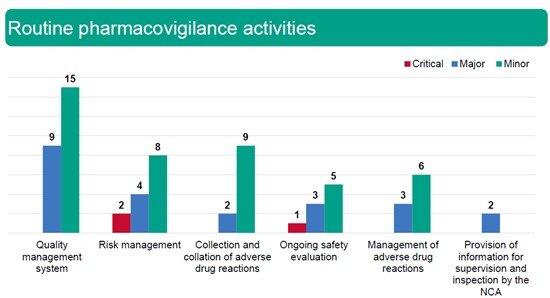 Figure 2: Inspection Findings of Routine Pharmacovigilance Activity
Source: Medicines and Healthcare products Regulatory Agency
During the reporting period of 2022/2023, four critical findings were issued and described below:
- Signal Management
- Maintenance of Reference Safety Information (RSI)
- Additional risk minimisation measures (aRMMs)
- Additional risk minimisation activities
The number of inspections from the previous reporting period (2021/2022) has reduced from 32 to 17, however, the mean number of inspection days has remained stable at around 9 days per inspection. The following topics remain the highest in terms of findings during the inspections:
- Risk Management
- Ongoing safety evaluation
- Quality management system
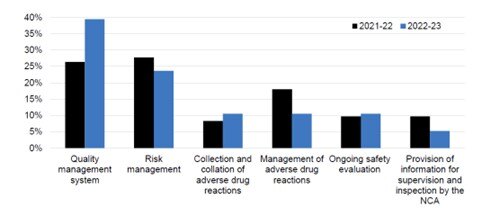 Figure 3: Inspection trends – distribution of major findings (as a percentage of total yearly findings) by topic
Source: Medicines and Healthcare products Regulatory Agency
Remote Compliance Assessment Pilot
A Remote Compliance Assessment was piloted in the GPvP team, and it is a new approach to following significant findings from previous inspections. Typically, a re-inspection would be conducted following a major or critical finding to ensure that the remediation activities were adequately implemented. However, this new approach replaces the need for a full re-inspection, under specific circumstances.
RCA
- Significant finding in one main topic
- Limited but targeted scope
- Always remote
- Document-based review of remediation is feasible.
Re-inspection
- Significant findings in multiple topics
- Remote inspection is not feasible
- Increased scope
- Remediation activities are complex or require interviews.
Where applicable, the RCA approach will be implemented to assess remediation of critical and major PV findings under certain circumstances. By using this approach, it is more time and cost effective for both the Inspector and MAH.
As there will be one focused topic to be considered, a full inspection plan, company presentation and pre-scheduled interviews will not be required. Instead, a focused RCA report form and a short introductory meeting will be held and a CAPA will only be raised if the finding is critical or major.
Conclusion
The GPvP team at the MHRA continue to improve their process of how inspections are conducted to ensure that it is time and cost-effective for all parties. It is crucial to ensure that the foundations, such as a QMS are in place for an MAH to establish and maintain the PV activities necessary for a fully compliant and risk-based approach for medicinal products.
Our Vigilance department at QbD Group will collaborate closely with you to understand your business needs and objectives and then offer specialised solutions to assist you in accomplishing them. With our expert guidance, you can confidently navigate pharmaceutical regulations.
Need regulatory support?
Trust our expertise to guide you through the regulatory process and achieve compliance
with ease.
Discover how we can help.
Stay ahead in life sciences
Keeping up with the fast-paced life sciences industry doesn’t have to be overwhelming.
Our newsletter delivers the latest insights, industry updates, and expert content directly to your inbox, helping you stay informed and make smarter decisions.

Discover more expert content
Webinar
From Strategy to First-in-Human: A Practical Roadmap for Biotech Development
Join our webinar to learn how to navigate nitrosamine risks and streamline your biotech development from early planning to first-in-human trials. Register now for expert insights.
Webinar
PMS & PMCF Compliance for Medical Devices Under EU MDR
Gain practical insights on PMS & PMCF compliance for medical devices under EU MDR. Watch our webinars on demand.
Case study
A Decade of Excellence: Linguistic Review Support for a Global Pharma Leader
QbD Group streamlines linguistic reviews for a leading pharma client, ensuring compliance and accuracy across 25 EU/EEA languages while meeting stringent regulatory deadlines.
Case study
Supporting Market Access to Essential Medicines in Europe
Support your entry into the European pharmaceutical market with expert regulatory guidance, ensuring compliance and access to essential medicines for patients. Contact us today for assistance.

Whitepaper
Medical Device Regulation (MDR) Checklist
Implement the Medical Device Regulation (MDR) with ease. Download our checklist of mandatory documents for MDR compliance.
Whitepaper
Regulatory Affairs for Pharma and Biotech
In this flyer, you will learn more about the regulatory services QbD Group provides for the pharmaceutical and biotechnology industries.
Whitepaper
Clinical Evaluation for Medical Devices under MDR
This whitepaper will guide you through crucial regulatory documents pertaining to the clinical evaluation process of your medical device. Download now!
Whitepaper
IVDR extension: what does this mean for you?
In this whitepaper, you will learn the impact of the IVDR transition extension and receive tips and strategies to navigate these regulatory changes.
Webinar
Drug-Device Combination Products and Article 117
Explore the regulatory complexities of Drug-Device Combination Products. Learn about Article 117 requirements, Notified Body Opinions, and more.
Whitepaper
Mobile health on the rise: exploring the regulatory landscape for reimbursement
This whitepaper will help you navigate the maze of the DTx regulatory environment, highlighting several important countries and regulations.
Whitepaper
Clinical investigations under MDR: key insights and practical experiences
This whitepaper covers clinical investigation authorization under MDR, the EU Legal Representative's role, and the MDR's impact on study conduct and post-market surveillance.
Case study
CE Mark Renewal for bioXtra Products: A Success Story
Discover how Lifestream Pharma successfully renewed the CE mark for their BioXtra Dry Mouth products under the EU MDR. Learn about the challenges faced, the approach taken for compliance, and the results achieved.
Webinar
Technical Documentation Essentials for Medical Device Software
Gain practical insights on crafting MDR-compliant technical documentation for Medical Device Software.

Webinar
Getting Started: Overcoming Initial Obstacles in Medical Device Software Development
Watch our webinar on demand and learn about regulatory obstacles, MDR, AI Act, and best practices for medical device software development and market entry.
Webinar
Mastering Clinical Performance Studies under IVDR
In the evolving regulatory landscape of IVDs, demonstrating clinical performance is more crucial and challenging than ever. Discover more in this webinar on demand.
Whitepaper
Clinical evidence for In Vitro Diagnostics under IVDR
In this whitepaper, we will guide you through the documents required for the demonstration of clinical evidence under the IVDR. Download now.
Webinar
Post-Market Surveillance Requirements for Medical Devices and IVDs
Learn about post-market surveillance requirements under MDR and IVDR regulations. Gain insights from industry experts in this on-demand webinar.
Whitepaper
The key role of Regulatory Affairs in the pharmaceutical industry: from drug development to commercialization
Discover the key role of Regulatory Affairs in pharma and how Regulatory Affairs teams support product lifecycle management in this whitepaper. Download now.

Webinar
3D Printing in the Healthcare World
Explore the challenges and opportunities of 3D printing in healthcare, featuring expert insights on regulations, pitfalls, and efficiency.
Webinar
IVDR Extension Explained: Tips & Strategies
Gain valuable insights on the IVDR extension and its impact on IVD manufacturers with our webinar on demand.
Whitepaper
Innovations in ATMP: state of the industry in 2024
This whitepaper explores the innovations in ATMP and solutions in detail, with a focus on the current landscape as of April 2024. Download now.
Case study
The journey toward IVDR compliance for the LVOne device
QbD Group helped Upfront Diagnostics gain entry into the MD/IVD industry by supporting them on their path to ISO 13485 certification.
Case study
Ensuring a smooth MDR transition for Oystershell's medical devices
The landscape of medical device regulations is constantly evolving, making it a challenge for companies to keep up. Our collaboration with Oystershell, which began in July 2023, highlights our expertise in transforming complex challenges into success stories, all with a touch of QbD Clinical magic.
Case study
Successful clinical evaluation supports rapid CE marking of Minze Health's Uroflowmeter
QbD Clinical assisted Minze Health, a urology-focused medical devices company, in obtaining CE marking under MDR through clinical evaluation.

Webinar
The Medical Device CE Certification Pathway Explained: From Classification to Conformity Assessment by the Notified Body
Learn about the Medical Device CE Certification Pathway, from classification to conformity assessment by a Notified Body. Join our webinar on demand.

Case study
Helping Relu to comply with changing regulations
QbD Group supported software company Relu with their expertise to comply with changing regulations.

Case study
Quickly bringing a new COVID-19 medical device to market
Obtaining the required government certifications for a medical device is always a challenge. Especially when the COVID clock is ticking.
Twitter Widget Iframe